Federal regulators cite insufficient survival evidence, concerns about China-heavy clinical trial data and additional safety risks, recommending another international study before approval.
The U.S. Food and Drug Administration has rejected Summit Therapeutics’ application for ivonescimab, an experimental treatment for certain patients with advanced lung cancer. The agency’s decision, announced November 15, 2026, followed a clinical trial that showed delayed cancer progression but failed to establish a statistically significant improvement in overall survival. A rejection letter made public on November 17 revealed additional concerns about trial design, patient safety and whether the results apply to American patients.
The decision blocks Summit’s attempt to introduce ivonescimab into the U.S. market under its current application. Regulators recommended another international clinical trial to establish whether the drug provides meaningful benefits that outweigh its risks. Summit maintains that its treatment has potential and plans to continue developing it for several types of cancer.
FDA Questions Evidence of Longer Survival
The application sought approval for ivonescimab combined with platinum-based chemotherapy in patients with locally advanced or metastatic non-small cell lung cancer. Eligible patients had tumors containing mutations in the EGFR gene and had previously received a targeted treatment known as an EGFR tyrosine kinase inhibitor. These patients can face limited treatment options when their cancer progresses.
Summit’s application relied primarily on results from the Phase 3 HARMONi clinical trial. The study compared ivonescimab plus chemotherapy against chemotherapy with a placebo, examining whether the experimental drug could delay worsening disease or extend patients’ lives. Both progression-free survival and overall survival were primary endpoints.
Patients receiving ivonescimab had a median progression-free survival of 6.8 months, compared with 4.4 months for patients receiving chemotherapy alone. The difference was statistically significant, indicating that the observed improvement was unlikely to be explained by chance under the trial’s statistical assumptions. However, the FDA considered the additional 2.4 months a modest benefit when weighed against the drug’s risks.
The study did not demonstrate a statistically significant improvement in overall survival. That finding does not prove ivonescimab cannot extend patients’ lives, but it means the submitted trial did not provide sufficiently convincing evidence of that benefit. Regulators also found no substantial difference in the overall response rate, which measures the proportion of patients whose tumors shrink by a defined amount.
Chinese Trial Data Raises Questions
The FDA’s newly published letter revealed substantial concerns about the geographic composition of the HARMONi trial. Although the study eventually included participants from North America and Europe, it originally enrolled patients exclusively in China. International enrollment did not begin until May 2023.
Approximately 60% of the trial’s participants were enrolled in China between January and November 2022. More importantly, 257 of the 275 progression-free survival events used in the primary analysis occurred among Chinese participants. That represented about 93% of the events supporting the primary efficacy finding.
Regulators said the concentration of those events prevented them from establishing whether the results could be reliably applied to American patients. Differences in patient characteristics, prior treatments and medical practices can affect how clinical findings translate across populations. The concern was about the representativeness of the evidence, rather than a determination that patients from China could not benefit from the treatment.
The FDA had raised the enrollment issue with Summit during discussions in March 2023 and again in May 2025. Those warnings preceded the final review, indicating that concerns about the study design were not new. The agency ultimately determined that expanding enrollment into Western countries had not resolved the limitations of the submitted evidence.
Additional Analysis Fails to Satisfy Regulators
Summit attempted to address the survival concerns through additional analyses based on longer patient follow-up. The company said these findings showed a favorable survival trend across Asian and Western patients. It reported an overall survival hazard ratio of 0.76 in its subsequent analysis, suggesting a possible reduction in the relative risk of death.
However, the FDA classified the post-hoc analyses as exploratory. Such analyses are performed or specified after researchers have examined trial data, making them less reliable for confirming a treatment benefit than prespecified comparisons. They can help generate new research questions but may not provide the evidence needed for regulatory approval.
Reviewers also found inconsistencies in the dates used to collect follow-up information from different regions. Some clinical sites in China stopped collecting data during 2025, earlier than sites in North America and Europe. That difference complicated comparisons between patient groups and further limited the agency’s confidence in the additional findings.
Safety Risks Influence FDA Decision
Safety was another important factor in the rejection. Ivonescimab is a bispecific antibody designed to target two biological pathways involved in cancer. It blocks PD-1, a protein that helps regulate immune responses, and VEGF, a protein involved in blood vessel growth.
The treatment is intended to help the immune system attack cancer while limiting the development of blood vessels that support tumor growth. However, interfering with these pathways can also cause harmful effects. FDA reviewers specifically cited serious immune-related complications and toxicities associated with VEGF inhibition.
The agency determined that the modest improvement in progression-free survival did not sufficiently outweigh the additional risks of combining ivonescimab with chemotherapy. Summit disagreed with that assessment, describing the treatment’s safety profile as acceptable and manageable. The company reported that 7.3% of patients receiving ivonescimab discontinued treatment because of treatment-related adverse events, compared with 5% in the control group.
Those figures reflect discontinuations rather than the full range of adverse reactions experienced during treatment. They also do not settle the broader question of whether the benefits justify additional toxicity. Regulators must evaluate the entire body of evidence when deciding whether a new medicine should become available.
Summit Plans to Continue Cancer Drug Development
Summit acquired development rights to ivonescimab in several international markets through a licensing agreement with China’s Akeso. The drug received its first marketing authorization in China in May 2024, but it remains investigational in the United States and Europe. Approval in one country does not automatically establish that a treatment meets another regulator’s standards.
The company had hoped ivonescimab could challenge established cancer immunotherapies, including Merck’s Keytruda. Earlier studies conducted in China produced encouraging results in different lung cancer settings. However, those findings cannot substitute for the evidence required to approve the specific treatment combination examined in HARMONi.
Summit President and Co-Chief Executive Officer Maky Zanganeh emphasized the importance of developing additional options for patients with difficult-to-treat cancers. Chairman and Co-Chief Executive Officer Robert Duggan also expressed disappointment with the decision while reaffirming the company’s commitment to ivonescimab. Summit continues to investigate the drug in lung, colorectal, bladder and kidney cancers.
The rejection may also influence expectations for other experimental treatments targeting both PD-1 and VEGF. Several pharmaceutical companies are developing therapies using similar approaches, although their trial designs, results and safety profiles differ. The FDA’s decision on ivonescimab does not determine whether those competing drugs will receive approval.
FDA Recommends a New International Trial
The FDA recommended that Summit conduct another adequately controlled, multiregional clinical trial designed to produce results applicable to patients in different countries. Regulators want statistically significant and clinically meaningful evidence of improved overall survival, progression-free survival or another appropriate measure of patient benefit. Any demonstrated improvement must also be sufficient to justify the treatment’s risks.
The agency advised Summit to discuss the study design and endpoints with regulators before beginning another trial. Such discussions could help the company avoid repeating problems involving enrollment, regional comparisons and the collection of survival information. A new trial would require additional time and resources, with no guarantee of eventual approval.
For patients with EGFR-mutated lung cancer, the immediate effect is that ivonescimab will not become an FDA-approved treatment through the rejected application. Its existing research programs will continue separately, and future regulatory decisions will depend on the evidence generated for each proposed use.
Summit’s next step is to determine whether it will pursue the new international trial recommended by the FDA. Until stronger evidence is available, regulators maintain that the proposed treatment’s benefits have not been shown to outweigh its risks for the patients covered by this application.
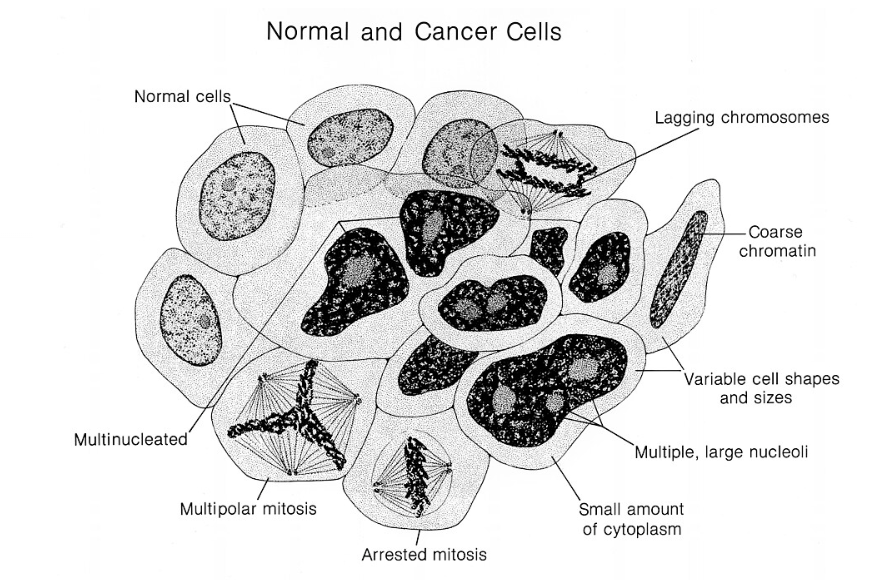
IMAGE: Author Pat Kenny This image was released by the National Cancer Institute CC0